Pharmaceutical Grade Enzyme for API Manufacturing
Troubleshoot GMP enzyme dosage, pH, temperature, QC, and supplier qualification for API manufacturing and drug formulation.
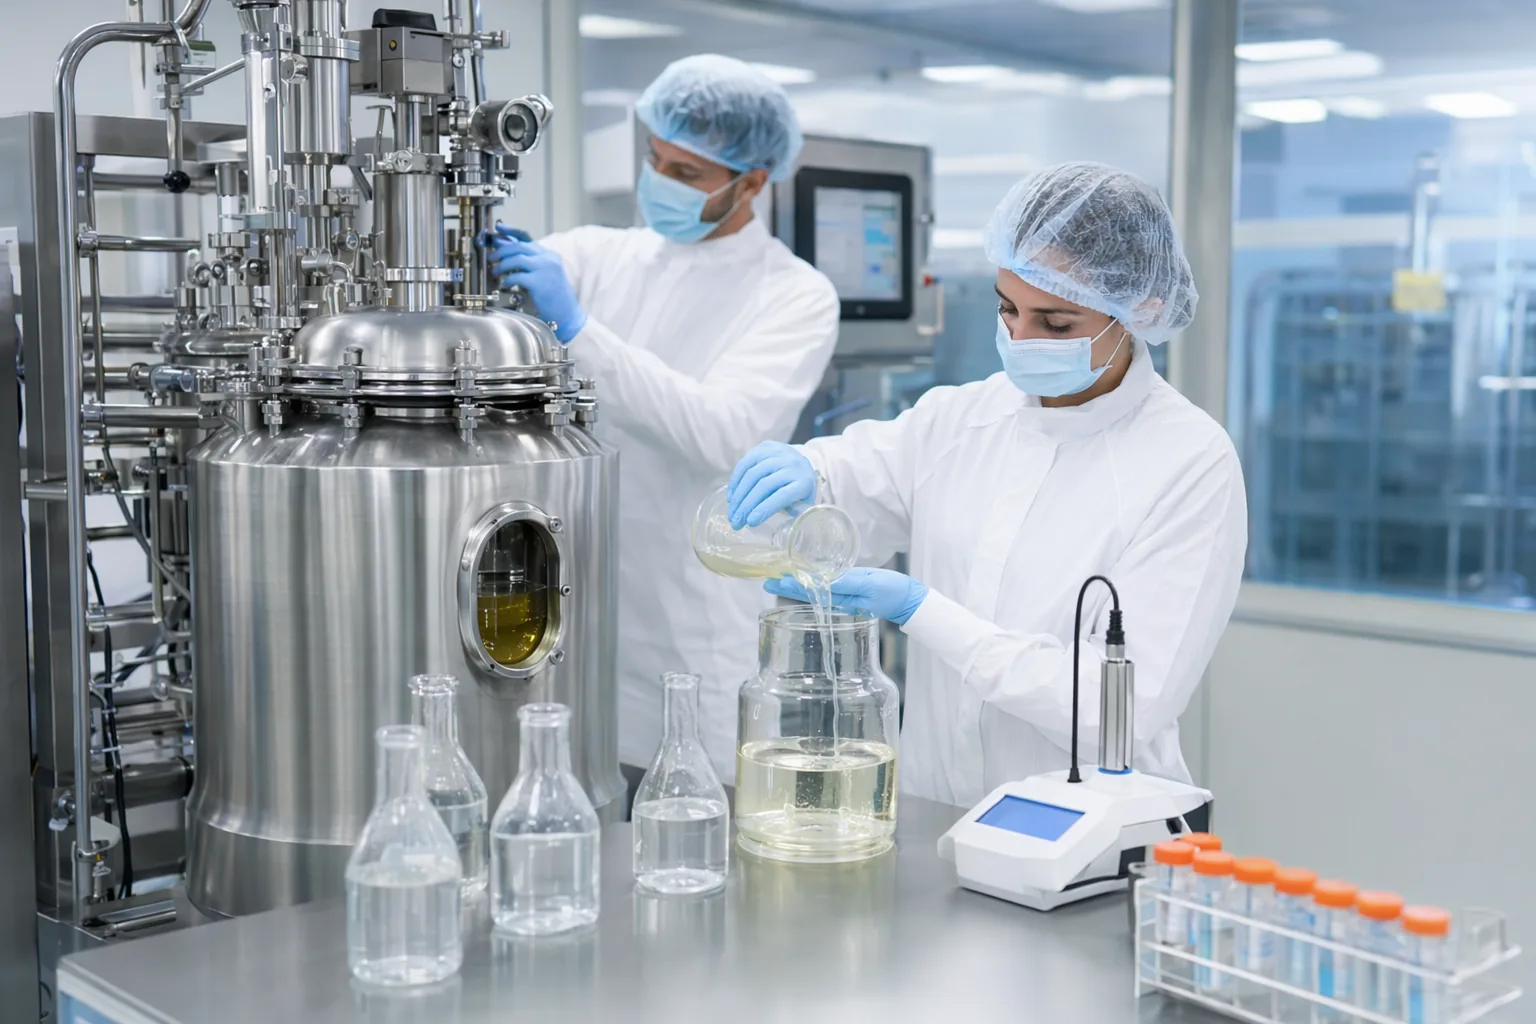
When an enzymatic API step loses conversion, forms new impurities or leaves residual activity, the cause is usually the process window, the activity definition or the removal step rather than the enzyme name on the label. This guide walks API manufacturers through troubleshooting, dosing by activity, pH and temperature control, release documents and pilot validation. To quote a grade for your route, send the enzyme, quantity and destination country.
Troubleshooting Enzyme Performance in API Manufacturing
A pharmaceutical-grade enzyme for API manufacturing must do more than show high catalog activity. In production, the enzyme must perform predictably in a defined solvent system, buffer, substrate concentration, pH range, and impurity profile. When conversion drops, first confirm the enzyme activity method on the COA, then compare it with your in-process assay rather than relying on mass dosage alone. Common causes include pH drift, denaturation at the charge temperature, substrate or product inhibition, inadequate mixing at high solids, and residual cleaning agents in equipment. For pharmaceutical, API manufacturing, and drug formulation workflows, EnzymePure supports a troubleshooting approach based on process data, pilot validation, and documented quality controls. This helps buyers judge an enzyme option by total process reliability instead of headline activity.
Verify enzyme activity units and assay conditions before scale-up. • Check pH and temperature at the actual reaction mass, not only the jacket. • Review impurities, solvents, salts, and preservatives for enzyme inhibition. • Confirm that residual enzyme removal or inactivation is validated.
Dosage Development: Start with Activity, Then Optimize Cost-in-Use
Effective dosage is normally established in activity units per kilogram of substrate, then translated into a practical weight or liquid addition rate for manufacturing. For early screening, set a low, mid and high dose from the activity stated for the lot and the supplier assay, rather than from weight alone. Highly selective transformations may need less, while viscous or high-solids reactions may require more to overcome mass transfer limitations. A capable supplier should help design a small DoE that tests dosage, substrate load, pH, temperature, hold time, and quench conditions. The best dosage is not always the fastest reaction; it is the point where conversion, impurity profile, filtration behavior, residual enzyme control, and cost-in-use meet the target process specification.
Run bench trials at low, mid, and high activity loading. • Track conversion, selectivity, impurity formation, and residual activity. • Calculate cost-in-use per kilogram of isolated API intermediate. • Avoid overdosing if it complicates downstream removal or validation.
pH, Temperature, and Hold-Time Controls
Most Pharmaceutical-Grade Enzymes have an operating window rather than a single ideal point. Start from the stated window of the grade (pH 2.0–8.5 depending on the enzyme, 25°C–40°C) and narrow it by enzyme class, buffer compatibility, substrate stability, and solvent tolerance. Some enzymes lose activity rapidly above their thermal threshold, while others show poor conversion at low temperature because substrate solubility or mixing becomes limiting. During troubleshooting, record the pH at reaction temperature and after substrate addition, because salts, acids, bases, and co-solvents can shift the effective environment. For drug formulation applications, processing temperatures may also be limited by excipient stability. An enzyme for pharmaceutical use should therefore be evaluated under the same pH, temperature, and hold-time conditions expected in the plant, not only under ideal laboratory assay conditions.
Screen pH in small steps around the expected optimum. • Screen temperature in small steps within stability limits. • Monitor pH drift during reaction and after quench. • Use validated inactivation, filtration, or removal steps where required.
QC Documentation: COA, TDS, SDS, and Release Checks
For regulated pharmaceutical processes, documentation is part of the enzyme specification. Buyers should request a COA that identifies lot number, activity, appearance, moisture or solids where relevant, microbiological limits when applicable, and any agreed impurity tests. The TDS should describe enzyme source, activity definition, recommended use conditions, storage, handling, and typical application guidance. The SDS should support safe handling, exposure control, spill response, and disposal planning. If a USP grade enzyme is requested, confirm the exact compendial expectation and whether it applies to the enzyme itself or to a related processing requirement. A pharmaceutical grade enzyme supplier for pharmaceutical projects should also support lot-to-lot comparison, retain samples, change notification expectations, and quality agreements where appropriate for the application and risk level.
Match the COA test methods to internal release requirements. • Confirm storage temperature, retest date, and shipping conditions. • Review allergen, animal-origin, and microbial control statements if relevant. • Document lot traceability through pilot and validation batches.
Supplier Qualification and Pilot Validation
Choosing a pharma enzyme supplier is a technical and quality decision. Qualification should include questionnaire review, documentation assessment, sample testing, change-control expectations, supply continuity, and technical responsiveness. A GMP enzyme requirement should be defined by process risk, regulatory filing strategy, and whether the enzyme remains in the final material or is removed during downstream purification. Before commercial purchase, run pilot validation at representative mixing, heat transfer, solids loading, solvent ratio, and filtration conditions. This is especially important for regulated API steps and drug formulation work. Compare candidates by performance data, documentation completeness, and cost-in-use so the selected enzyme is practical for scale-up and supplier qualification.
Qualify suppliers before locking the regulatory or process control strategy. • Use pilot batches to confirm kinetics, impurity profile, and removal steps. • Assess backup supply options for business continuity. • Include technical support responsiveness in supplier scoring.
Define the Failure Mode Before Changing the Enzyme
Troubleshooting should diagnose the process variable before anyone quotes a different enzyme. In API manufacturing, enzymes may be used for selective biocatalysis, impurity reduction, resolution, deprotection, or mild processing steps. In drug formulation, the priority may be excipient modification, controlled processing, or removal of a process aid within a defined specification. The first troubleshooting step is to define the failure mode: low conversion, side-product formation, slow reaction rate, viscosity increase, filtration difficulty, residual activity, or batch-to-batch variability. Share substrate, solvent or buffer system, target pH, temperature, hold time, acceptable residuals, and downstream purification limits under confidentiality. Expect a supplier to answer with a grade recommendation and its documents, and plan pilot-scale validation on your own equipment, rather than relying on unverifiable performance claims.
Define the process objective and acceptance criteria. • Separate enzyme activity issues from mixing, mass transfer, or raw material variability. • Check whether the application is API manufacturing, drug formulation, or an ancillary processing step.
Technical Buying Checklist
Confirm the activity definition and assay conditions for the lot match your in-process assay. • Measure pH and temperature in the reaction mass, not only at the jacket. • Test a low, mid and high activity-based dose and track conversion, selectivity and impurities. • Check solvents, salts and cleaning residues for inhibition. • Validate the quench, inactivation or removal step before scale-up.
Request a Quote for an API Route
Tell us the enzyme class, the quantity per campaign and the destination country, and describe the reaction step (substrate, solvent or buffer, target conditions). Grades are supplied as white to off-white lyophilized powder from 100 g; price and lead time are confirmed per order.
Pharmaceutical-Grade Enzymes for Research & Industry
Need Pharmaceutical-Grade Enzymes for your lab or production process?
ISO 9001 certified · Food-grade & research-grade · Ships to 80+ countries
Frequently Asked Questions
How do I select an enzyme supplier for API manufacturing?
Start with technical fit, then qualify the quality system. Ask for COA, TDS, SDS, activity method, traceability, storage conditions, and change notification expectations. Test multiple lots when possible. The supplier should also help troubleshoot pH, temperature, dosage, inhibition, downstream removal, and cost-in-use before you move into validation or routine purchasing.
What dosage should we use for an enzyme in drug formulation?
Dosage depends on enzyme activity, substrate or excipient load, water activity, pH, temperature, hold time, and removal or inactivation requirements. Early trials usually compare a low, mid and high activity-based dose. In drug formulation, choose the lowest robust dose that meets performance targets without creating downstream, stability, or residual enzyme concerns.
What pH and temperature ranges are safe starting points?
The grades are specified for pH 2.0–8.5 (enzyme-dependent) and 25°C–40°C; the working range inside that window depends on enzyme class and process chemistry. Check the TDS first, then run bench trials in small increments. For a drug formulation process, also consider excipient stability, microbial control strategy, and whether temperature affects viscosity or mixing.
Do we need a GMP enzyme for every API process?
Not always. The need for a GMP enzyme depends on its role, regulatory strategy, removal capability, residual risk, and whether it contacts material that will become part of the final API or drug product. Define requirements with quality and regulatory teams. Even when full GMP is not required, robust documentation, traceability, COA review, and supplier qualification are still important.
How should cost-in-use be compared between enzyme suppliers?
Do not compare enzyme price per kilogram alone. Compare activity-adjusted dosage, conversion rate, yield, impurity reduction, batch time, downstream filtration, inactivation needs, residual testing, waste, and supply reliability. A higher-priced enzyme may reduce total cost if it improves selectivity, shortens hold time, or simplifies purification at manufacturing scale.
Turn This Guide Into a Supplier Brief
Request a technical review for your API or drug formulation enzyme process. See our application page for Pharmaceutical-Grade Enzymes for Controlled GMP Production for specs, MOQ, and a free 50 g sample.
Contact Us to Contributehello@mail.enzymepure.com